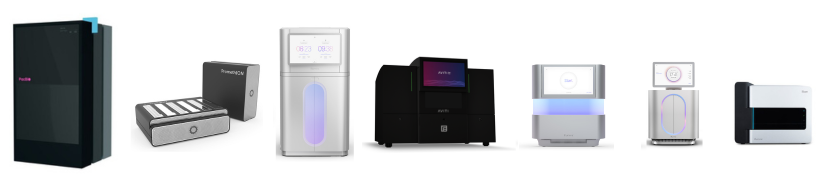
NGI is one of the largest technical platforms at SciLifeLab. We provide access to technology for sequencing, genotyping and associated bioinformatics support to researchers based in Sweden.
NGI OpenLab: A New Hub for Collaborative Genomics!
We're thrilled to announce the official launch of NGI OpenLab, an innovative space designed to empower genomics research. The lab provides direct access to equipment for quality control (QC), library preparation and a walk-up sequencer for on-the-go sequencing needs.
At NGI, we continuously monitor our sequencing output to ensure the highest standards of data quality. Over the past year, our team has been actively investigating intermittent data quality observations on our NextSeq 2000 platform. Below is a detailed overview of the issue, our interactions with Illumina, and the steps we are taking to resolve it.
NGI now offers the Watchmaker DNA Library Prep Kit with Fragmentation, providing both PCR-free and PCR-based workflows with lower DNA input requirements.
Want to stay in touch? Join our newsletter!
All the latest updates from NGI direct to your inbox!
A comprehensive comparison of RNA-Seq-based transcriptome analysis from reads to differential gene expression and cross-comparison with microarrays: a case study in Saccharomyces cerevisiae.
I Nookaew, M Papini, N Pornputtapong, G Scalcinati, L Fagerberg, M Uhlén, J Nielsen
Nucleic Acids Res., 40 (20) 1362-4962 (2012)
RNA-seq, has recently become an attractive method of choice in the studies of transcriptomes, promising several advantages compared with microarrays. In this study, we sought to assess the contribution of the different analytical steps involved in the analysis of RNA-seq data generated with the Illumina platform, and to perform a cross-platform comparison based on the results obtained through Affymetrix microarray. As a case study for our work we, used the Saccharomyces cerevisiae strain CEN.PK 113-7D, grown under two different conditions (batch and chemostat). Here, we asses the influence of genetic variation on the estimation of gene expression level using three different aligners for read-mapping (Gsnap, Stampy and TopHat) on S288c genome, the capabilities of five different statistical methods to detect differential gene expression (baySeq, Cuffdiff, DESeq, edgeR and NOISeq) and we explored the consistency between RNA-seq analysis using reference genome and de novo assembly approach. High reproducibility among biological replicates (correlation≥0.99) and high consistency between the two platforms for analysis of gene expression levels (correlation≥0.91) are reported. The results from differential gene expression identification derived from the different statistical methods, as well as their integrated analysis results based on gene ontology annotation are in good agreement. Overall, our study provides a useful and comprehensive comparison between the two platforms (RNA-seq and microrrays) for gene expression analysis and addresses the contribution of the different steps involved in the analysis of RNA-seq data.
EM Toledo, S Yang, D Gyllborg, KE van Wijk, I Sinha, M Varas-Godoy, CL Grigsby, P Lönnerberg, S Islam, KR Steffensen, S Linnarsson, E Arenas
Cell Rep, 31 (5) 2211-1247 (2020)
Liver X receptors (LXRs) and their ligands are potent regulators of midbrain dopaminergic (mDA) neurogenesis and differentiation. However, the molecular mechanisms by which LXRs control these functions remain to be elucidated. Here, we perform a combined transcriptome and chromatin immunoprecipitation sequencing (ChIP-seq) analysis of midbrain cells after LXR activation, followed by bioinformatic analysis to elucidate the transcriptional networks controlling mDA neurogenesis. Our results identify the basic helix-loop-helix transcription factor sterol regulatory element binding protein 1 (SREBP1) as part of a cluster of proneural transcription factors in radial glia and as a regulator of transcription factors controlling mDA neurogenesis, such as Foxa2. Moreover, loss- and gain-of-function experiments in vitro and in vivo demonstrate that Srebf1 is both required and sufficient for mDA neurogenesis. Our data, thus, identify Srebf1 as a central player in mDA neurogenesis.
Consequences of past climate change and recent human persecution on mitogenomic diversity in the arctic fox.
P Larsson, J von Seth, IJ Hagen, A Götherström, S Androsov, M Germonpré, N Bergfeldt, S Fedorov, NE Eide, N Sokolova, D Berteaux, A Angerbjörn, Ø Flagstad, V Plotnikov, K Norén, D Díez-Del-Molino, N Dussex, DWG Stanton, L Dalén
Ancient DNA provides a powerful means to investigate the timing, rate and extent of population declines caused by extrinsic factors, such as past climate change and human activities. One species probably affected by both these factors is the arctic fox, which had a large distribution during the last glaciation that subsequently contracted at the start of the Holocene. More recently, the arctic fox population in Scandinavia went through a demographic bottleneck owing to human persecution. To investigate the consequences of these processes, we generated mitogenome sequences from a temporal dataset comprising Pleistocene, historical and modern arctic fox samples. We found no evidence that Pleistocene populations in mid-latitude Europe or Russia contributed to the present-day gene pool of the Scandinavian population, suggesting that postglacial climate warming led to local population extinctions. Furthermore, during the twentieth-century bottleneck in Scandinavia, at least half of the mitogenome haplotypes were lost, consistent with a 20-fold reduction in female effective population size. In conclusion, these results suggest that the arctic fox in mainland Western Europe has lost genetic diversity as a result of both past climate change and human persecution. Consequently, it might be particularly vulnerable to the future challenges posed by climate change. This article is part of a discussion meeting issue 'The past is a foreign country: how much can the fossil record actually inform conservation?'
Comparison of spleen transcriptomes of two wild rodent species reveals differences in the immune response against Borrelia afzelii.
X Zhong, M Lundberg, L Råberg
Ecol Evol, 10 (13) 2045-7758 (2020)
Different host species often differ considerably in susceptibility to a given pathogen, but the causes of such differences are rarely known. The natural hosts of the tick-transmitted bacterium Borrelia afzelii, which is one of causative agents of Lyme borreliosis in humans, include a variety of small mammals like voles and mice. Previous studies have shown that B. afzelii-infected bank voles (Myodes glareolus) have about ten times higher bacterial load than infected yellow-necked mice (Apodemus flavicollis), indicating that these two species differ in resistance. In this study, we compared the immune response to B. afzelii infection in these host species by using RNA sequencing to quantify gene expression in spleen. Gene set enrichment analysis (GSEA) showed that several immune pathways were down-regulated in infected animals in both bank voles and yellow-necked mice. Moreover, IFNα response was up-regulated in B. afzelii-infected yellow-necked mice, while IL6 signaling and the complement pathway were down-regulated in infected bank voles; differences in regulation of these three pathways between bank voles and yellow-necked mice could thus contribute to the difference in resistance to B. afzelii between the species. This study provides knowledge of gene expression induced by a zoonotic pathogen in its natural host, and possible species-specific regulation of immune responses associated with resistance.
Patients with systemic lupus erythematosus (SLE) have an increased bisphenol A methylation score linked to SLE risk genes and selected clinical subphenotypes.
H Vestin, N Oparina, ML Eloranta, M Frodlund, I Gunnarsson, C Sjöwall, E Svenungsson, L Rönnblom, J Imgenberg-Kreuz, D Leonard
RMD Open, 11 (3) 2056-5933 (2025)
Bisphenol A (BPA), a xenoestrogen that can alter DNA methylation status, has been implicated in the pathogenesis of systemic lupus erythematosus (SLE). This study aimed to investigate whether methylation changes at BPA-sensitive 5'-C-phosphate-G-3' (CpG) sites are associated with SLE and clinical subphenotypes.
A discovery cohort (n=747) and a replication cohort (n=388) including Swedish patients with SLE and healthy controls were investigated using the Illumina HM450k bead chip. BPA-sensitive CpG sites were selected if differentially methylated in ≥2 of 7 BPA exposure studies and supported by cell line data. A BPAAll score including 19 CpGs and a BPASLE score based on three CpG sites co-localised in the genome with SLE risk loci were calculated for each individual, analysed for associations with clinical data and then compared with publicly available transcriptomic data from BPA-treated cells.
Patients with SLE had significantly higher BPASLE score than controls in the discovery (OR 1.34, p=4.6×10⁻¹³), replication (OR 1.28, p=1.1×10⁻⁵) and meta-analysis (OR 1.32, p=3.3×10⁻¹⁷). Higher BPAAll score was associated with SLE in the discovery cohort (OR 1.05, p=2.3×10⁻³) but not in the replication cohort (OR 1.04, p=0.12) with a significant difference in the meta-analysis (OR 1.05, p=7.0×10⁻⁴). Both scores were associated with prednisolone treatment (p<0.001), and the BPASLE score was associated with serositis and autoantibodies (p<0.05). Transcriptomic analysis of BPA-treated cells revealed enrichment in pathways such as interferon and mitogen-activated protein kinase signalling.
Our findings reveal a novel association between BPA exposure and DNA methylation changes in SLE, with potential implications for the regulation of immune-related gene expression.
miR-128 regulates non-myocyte hyperplasia, deposition of extracellular matrix and Islet1 expression during newt cardiac regeneration.
N Witman, J Heigwer, B Thaler, WO Lui, JI Morrison
Dev. Biol., 383 (2) 1095-564X (2013)
Cardiovascular disease is a global scourge to society, with novel therapeutic approaches required in order to alleviate the suffering caused by sustained cardiac damage. MicroRNAs (miRNAs) are being touted as one such approach in the fight against heart disease, acting as possible post-transcriptional molecular triggers responsible for invoking cardiac regeneration. To further ones understanding of miRNAs and cardiac regeneration, it is prudent to learn from organisms that can intrinsically regenerate their hearts following injury. Using the red-spotted newt, an adult chordate capable of cardiac regeneration, we decided to delve deeper into the role miRNAs play during this process. RNA isolated from regenerating newt heart samples, was used in a microarray screen, to identify significantly expressed candidate miRNAs during newt cardiac regeneration. We performed quantitative qPCR analysis on several conserved miRNAs and found one in particular, miR-128, to be significantly elevated when cardiac hyperplasia is at its peak following injury. In-situ hybridisation techniques revealed a localised expression pattern for miR-128 in the cardiomyocytes and non-cardiomyocytes in close proximity to the regeneration zone and in vivo knockdown studies revealed a regulatory role for miR-128 in proliferating non-cardiomyocyte populations and extracellular matrix deposition. Finally, 3'UTR reporter assays revealed Islet1 as a biological target for miR-128, which was confirmed further through in vivo Islet1 transcriptional and translational expression analysis in regenerating newt hearts. From these studies we conclude that miR-128 regulates both cardiac hyperplasia and Islet1 expression during newt heart regeneration and that this information could be translated into future mammalian cardiac studies.
Whole genome case-control study of central nervous system toxicity due to antimicrobial drugs.
J Ås, I Bertulyte, N Norgren, A Johansson, N Eriksson, H Green, M Wadelius, P Hallberg
A genetic predisposition to central nervous system (CNS) toxicity induced by antimicrobial drugs (antibiotics, antivirals, antifungals, and antiparasitic drugs) has been suspected. Whole genome sequencing of 66 cases and 833 controls was performed to investigate whether antimicrobial drug-induced CNS toxicity was associated with genetic variation. The primary objective was to test whether antimicrobial-induced CNS toxicity was associated with seventeen efflux transporters at the blood-brain barrier. In this study, variants or structural elements in efflux transporters were not significantly associated with CNS toxicity. Secondary objectives were to test whether antimicrobial-induced CNS toxicity was associated with genes over the whole genome, with HLA, or with structural genetic variation. Uncommon variants in and close to three genes were significantly associated with CNS toxicity according to a sequence kernel association test combined with an optimal unified test (SKAT-O). These genes were LCP1 (q = 0.013), RETSAT (q = 0.013) and SFMBT2 (q = 0.035). Two variants were driving the LCP1 association: rs6561297 (p = 1.15x10⁻⁶, OR: 4.60 [95% CI: 2.51-8.46]) and the regulatory variant rs10492451 (p = 1.15x10⁻⁶, OR: 4.60 [95% CI: 2.51-8.46]). No common genetic variant, HLA-type or structural variation was associated with CNS toxicity. In conclusion, CNS toxicity due to antimicrobial drugs was associated with uncommon variants in LCP1, RETSAT and SFMBT2.
Last Updated: 7th July 2026
This website uses cookies to improve your experience. AcceptRead More
Privacy & Cookies Policy
Privacy Overview
This website uses cookies to improve your experience while you navigate through the website. Out of these, the cookies that are categorized as necessary are stored on your browser as they are essential for the working of basic functionalities of the website. We also use third-party cookies that help us analyze and understand how you use this website. These cookies will be stored in your browser only with your consent. You also have the option to opt-out of these cookies. But opting out of some of these cookies may affect your browsing experience.
Necessary cookies are absolutely essential for the website to function properly. This category only includes cookies that ensures basic functionalities and security features of the website. These cookies do not store any personal information.
Any cookies that may not be particularly necessary for the website to function and is used specifically to collect user personal data via analytics, ads, other embedded contents are termed as non-necessary cookies. It is mandatory to procure user consent prior to running these cookies on your website.